
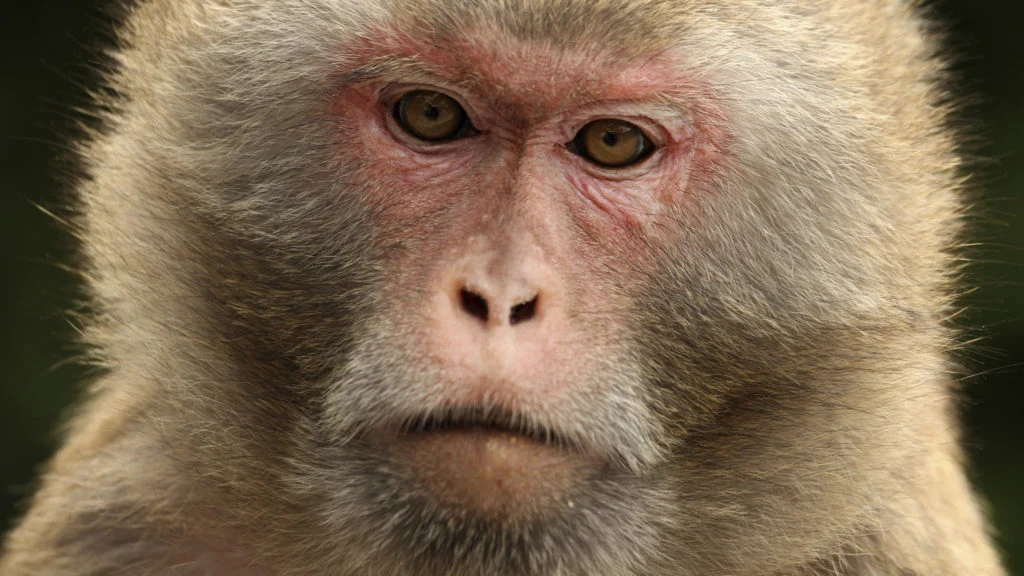
In a decisive move to reshape the landscape of pharmaceutical research, the Department of Health and Human Services (HHS) and the Food and Drug Administration (FDA) have officially finalized a new regulatory framework designed to expedite the transition toward human-centric testing methods. The final rule, published on September 22, 2026, marks the second major policy shift this year aimed at reducing the medical industry’s reliance on animal models, signaling a pivot toward more sophisticated, nonclinical technologies such as organ-on-a-chip platforms, computer modeling, and advanced cell cultures.
The core of the regulatory update is semantic yet profound: the FDA is officially replacing legacy terminology within its guidelines, swapping phrases like "animal tests" and "animal studies" for the more inclusive "nonclinical tests" and "nonclinical studies." While the change may appear purely administrative, it establishes a formal legal pathway for researchers to bypass traditional animal testing requirements when alternative methods are deemed scientifically appropriate to determine the safety and efficacy of new drugs and biologics.
A Chronology of Regulatory Evolution
The movement to replace animal testing has been gaining momentum for over a decade, driven by technological breakthroughs and mounting pressure from both animal welfare organizations and the scientific community.
- 2010-2015: Initial investment in "organ-on-a-chip" technology begins to show viability, allowing scientists to mimic human physiological responses on microchips.
- 2022: The FDA Modernization Act 2.0 is signed into law, granting the agency the authority to approve drugs that have been tested using alternative methods, provided they are sufficiently robust, without strictly requiring animal data.
- Early 2026: HHS announces the first set of strategic initiatives focused on incentivizing the adoption of human-based research models across federally funded projects.
- September 22, 2026: The FDA issues its final rule, standardizing terminology to facilitate the integration of non-animal methodologies into the formal regulatory filing process.
The Scientific Case for Human-Based Research
For decades, the standard pharmaceutical development pipeline required animal testing to evaluate toxicity and safety before clinical trials in humans could commence. However, critics of this model have long pointed to a significant translational gap. Biological differences between species often lead to misleading results; a compound that proves safe in a rodent model may trigger unforeseen side effects in humans, or conversely, a promising therapy may be discarded because it performed poorly in animals despite potential human utility.
Data from the National Institutes of Health (NIH) indicates that approximately 90% of drugs that pass animal trials ultimately fail in human clinical trials, often due to a lack of safety or efficacy. By integrating human-based platforms—such as microphysiological systems, three-dimensional cell cultures, and artificial intelligence-driven predictive modeling—researchers hope to lower this failure rate.

These modern tools allow for the testing of drugs on human tissue that carries specific genetic markers or disease profiles, providing a level of precision that animal models simply cannot replicate. Furthermore, these methods often offer faster turnaround times and lower costs, potentially reducing the overall burden of the drug development lifecycle.
Official Stance and Administrative Strategy
The administration’s push is being framed as a necessary modernization of a regulatory system that has remained static for nearly half a century. Health Secretary Robert F. Kennedy Jr. underscored the significance of this shift in a formal statement following the announcement.
"We are moving HHS toward a new era of biomedical research that puts human biology at the center of science," Kennedy said. "We are modernizing outdated regulations, investing in human-based technologies, and breaking down barriers that have kept researchers dependent on animal models when better tools are available."
The HHS strategy involves not only changing regulations but also reallocating research grants toward high-throughput screening technologies. The administration contends that by aligning FDA terminology with current scientific realities, they are providing clear guidance to pharmaceutical companies, effectively removing the "regulatory fear" that previously prevented firms from choosing advanced nonclinical methods over traditional animal studies.
Industry Implications and Market Impact
The pharmaceutical industry has reacted with cautious optimism. For large-scale drug developers, the transition represents both an opportunity and a challenge. On one hand, the ability to utilize faster, more accurate testing methods could significantly shorten the time-to-market for new drugs. On the other, the industry must invest heavily in the infrastructure required to adopt these new technologies, including staff training and the implementation of proprietary data-analysis tools.
"The industry is supportive of innovation, but the transition must be handled with scientific rigor," noted a spokesperson for a major biotechnology trade association. "The primary concern remains ensuring that regulators—globally—will accept these new forms of data as sufficient evidence of safety, especially when seeking approval for novel therapies."

There is also a growing concern regarding international harmonization. Because clinical trials and drug approvals are often global processes, the FDA’s new rules must eventually align with the European Medicines Agency (EMA) and other international bodies. If the FDA accepts nonclinical data that other regulators refuse, manufacturers may find themselves stuck in a "double-testing" cycle, negating the time and cost savings the rule aims to create.
Analyzing the Long-Term Trajectory
The broader impact of this rule change extends beyond the pharmaceutical sector. It influences the academic research pipeline, where the next generation of scientists is being trained. As the FDA sets the standard for "appropriate" nonclinical testing, academic institutions will likely shift their curriculum and laboratory funding to reflect these priorities.
However, a critical analysis suggests that this is not an overnight phase-out. Animal research remains a requirement for specific complex biological questions, particularly those involving whole-organism interactions, such as systemic immune responses or complex neurobiological pathways. The FDA’s language, which includes the qualifier "when appropriate," suggests that the agency intends to transition slowly, using a hybrid model for the foreseeable future.
The effectiveness of this policy will ultimately be measured by the number of successful drug applications (NDAs) that utilize these new, non-animal methodologies over the coming years. If the FDA succeeds in creating a clear, predictable, and scientifically sound path for these submissions, it could mark the most significant change in pharmaceutical safety testing since the 1960s.
Conclusion
The September 22 rule is a clear signal that the FDA is prioritizing technological modernization over traditional proceduralism. While the transition will be gradual and will require significant investment from both public and private sectors, the shift toward human-based testing models represents a fundamental change in the philosophy of medicine. By centering human biology in the earliest stages of the drug development process, the administration aims to create a more efficient, safer, and more scientifically accurate future for medical innovation.
As the implementation phase begins, the scientific community will be watching closely to see how the FDA defines the threshold for "appropriateness" in nonclinical testing. For now, the regulatory foundation has been laid, setting the stage for a, perhaps, less animal-dependent future in the search for the next generation of life-saving therapies.




Leave a Reply